Sequana documentation#
0.24.2
SEQUANA#
)


- How to cite:
Citations are important for us to carry on developments. For Sequana library (including the pipelines), please use
Cokelaer et al, (2017), 'Sequana': a Set of Snakemake NGS pipelines, Journal of Open Source Software, 2(16), 352, JOSS DOI doi:10.21105/joss.00352
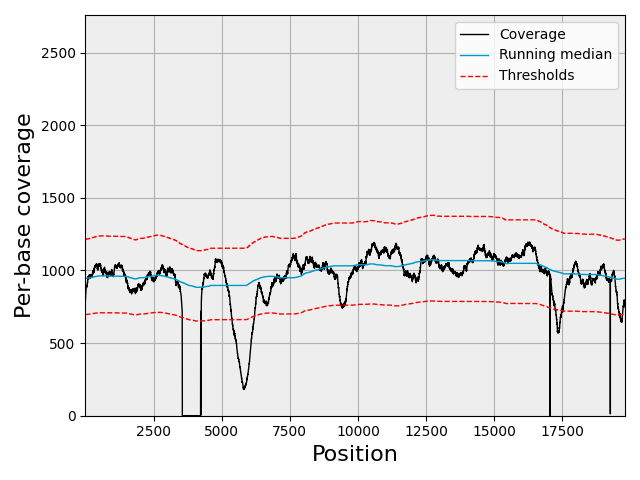
For the genome coverage tool (sequana_coverage): Desvillechabrol et al, 2018: detection and characterization of genomic variations using running median and mixture models. GigaScience, 7(12), 2018. https://doi.org/10.1093/gigascience/giy110
For Sequanix: Desvillechabrol et al. Sequanix: A Dynamic Graphical Interface for Snakemake Workflows Bioinformatics, bty034, https://doi.org/10.1093/bioinformatics/bty034 Also available on bioRxiv (DOI: https://doi.org/10.1101/162701)
🔧 Overview and Installation#
Sequana is a Python library dedicated to bioinformatics. It is also a project that includes a set of pipelines related to NGS (new generation sequencing) including quality control, variant calling, coverage, taxonomy, transcriptomics. We also ship Sequanix, a graphical user interface for Snakemake pipelines.
Pipelines and related projects#
Here is a non-exhaustive list of tools and pipelines from the project, with users and developers audience.
name/github |
description |
Latest Pypi version |
Test passing |
apptainers |
|---|---|---|---|---|
Create and Manage Sequana pipeline |

|

|
Not required |
|
Set of wrappers to build pipelines |

|

|
Not required |
|
Demultiplex your raw data |

|

|
License restriction |
|
denovo sequencing data |

|

|

|
|
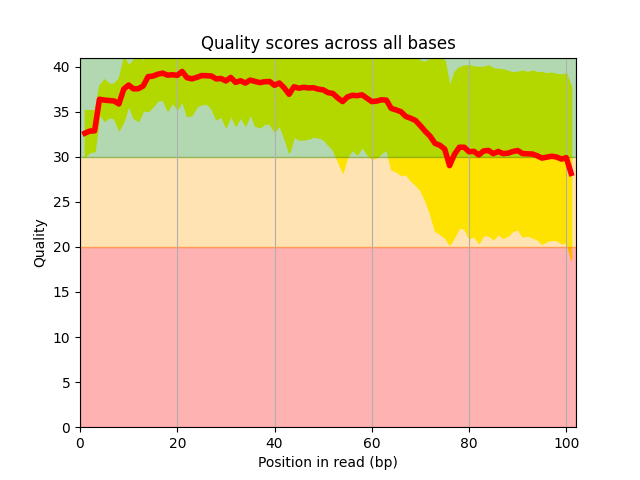
Get Sequencing Quality control |

|

|

|
|
Map sequences on target genome |

|

|

|
|
Map sequences on target genome |

|

|

|
|
Merge barcoded (or unbarcoded) nanopore fastq and reporting |

|

|

|
|
Pacbio quality control |

|

|

|
|
Find ribosomal content |

|

|

|
|
RNA-seq analysis |

|

|

|
|
Variant Calling |

|

|

|
|
Coverage (mapping) |

|

|

|
|
Long read Amplicon Analysis |

|

|

|
|
reverse complement of sequence data |

|

|

|
|
downsample sequencing data |

|

|
Not required |
|
remove/select reads mapping a reference |
|

|
name/github |
description |
Latest Pypi version |
Test passing |
|---|---|---|---|
Find repeats |

|

|
|
Taxonomy analysis |

|

|
Please see the documentation for an up-to-date status and documentation.
Contributors#
Maintaining Sequana would not have been possible without users and contributors. Each contribution has been an encouragement to pursue this project. Thanks to all:
Changelog :memo:#
Version |
Description |
|---|---|
0.24.2 |
|
0.24.1 |
|
0.24.0 |
|
0.23.0 |
|
0.22.0 |
|
0.21.2 |
|
0.21.1 |
|
0.21.0 |
|
0.20.0 |
|
0.19.6 |
|
0.19.5 |
|
0.19.4 |
|
0.19.3 |
|
0.19.2 |
|
0.19.1 |
|
0.19.0 |
|
0.18.0 |
|
0.17.3 |
|
0.17.2 |
|
0.17.1 |
|
0.17.0 |
|
0.16.9 |
|
0.16.8 |
|
0.16.7 |
|
0.16.6 |
|
0.16.5 |
|
0.16.4 |
|
0.16.3 |
|
0.16.2 |
|
0.16.1 |
|
0.16.0 |
|
0.15.4 |
|
0.15.3 |
|
0.15.2 |
|
0.15.1 |
|
0.15.0 |
|
0.14.6 |
|
0.14.5 |
|
0.14.4 |
|
0.14.3 |
|
0.14.2 |
|
0.14.1 |
|
0.14.0 |
|
0.13.X |
|
0.12.X |
|
Any :question: Feel free to [open an issue](sequana/sequana#issues)
What is Sequana?#
Sequana is a versatile bioinformatics tool that provides:
A Python library dedicated to NGS analysis (file-format wrappers, plots, coverage, taxonomy, enrichment, RNA-seq, …).
A family of pipelines based on Snakemake, each living in its own GitHub repository.
- A set of standalone applications:
sequana_coverage — genome coverage analysis with confidence intervals.
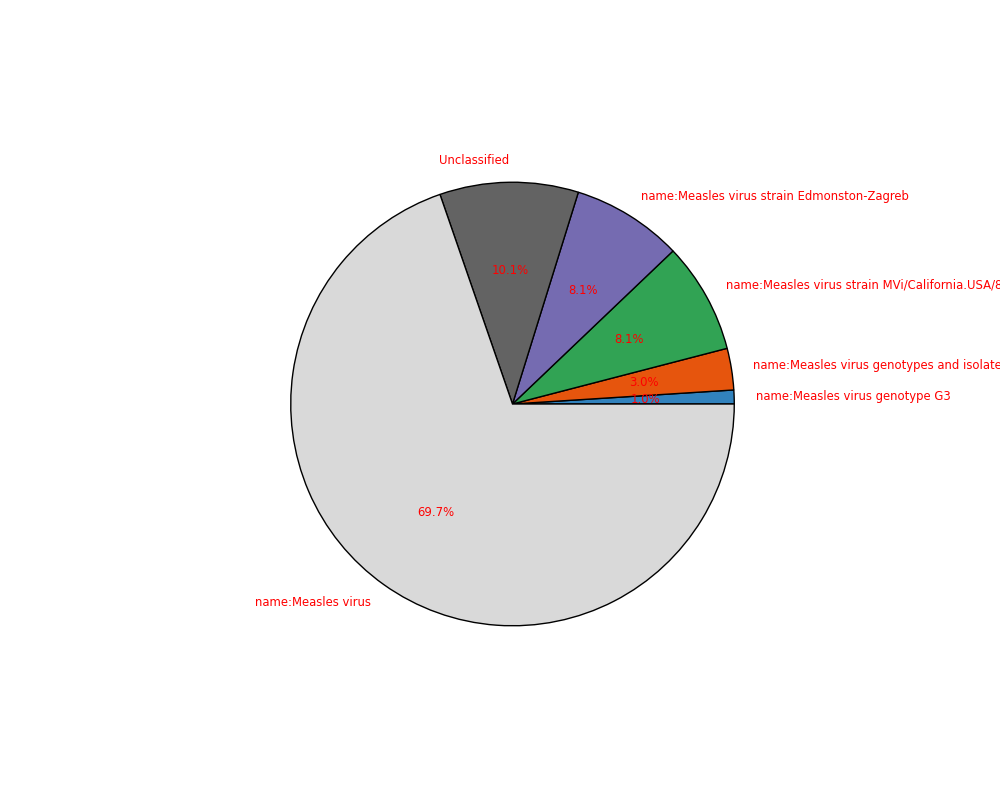
sequana_taxonomy — quick taxonomy on a FastQ file (Kraken + Krona).
~30 other utility sub-commands grouped under the top-level
sequanaCLI (see Applications (standalone)).
Pipelines cover NGS quality control, variant calling, coverage analysis, taxonomy, de-novo assembly, RNA-seq, variant calling and more — see the Pipelines catalogue.
Sequana can be used either as a Python library (developers, library users) or via its pipelines and standalones (end users). To join the project, please let us know on github.
Quick start#
Install Sequana and run a quality-control pipeline in 2 minutes:
pip install sequana_fastqc
sequana_fastqc --input-directory . --execute
Or use the Python library to read and analyze genomic data:
from sequana import FastA, SequanaCoverage
fasta = FastA("genome.fasta")
cov = SequanaCoverage("coverage.bed")
cov[0].plot_coverage()
Full guides below.
Choosing your entry point#
Not sure where to start? This table helps:
What you want to do |
Use this |
|---|---|
Run a pre-built NGS pipeline (fastqc, variant calling, RNA-seq, etc.) |
|
Use Python to read/analyze sequences, coverage, or variants |
|
Find a quick helper tool (rename files, filter GFF, count bases) |
|
See working code examples |
Installation
pip install sequana
Library User Guide
Use Sequana from Python to read, analyze, and visualize genomic data
Tutorial
Step-by-step guides for common workflows and pipelines
Code Examples
Runnable Python examples from the gallery
NGS Pipelines
Browse the Snakemake pipelines catalogue
CLI Reference
The sequana command-line tools (30+ sub-commands)
Pipeline users
Library users
Reference & developers
- Developer guide
- Applications (standalone)
- Sequanix Tutorial
- Wrappers
- bowtie2/align
- bowtie2/build
- add_read_group
- bam_coverage
- bcl2fastq
- blast
- busco
- bz2_to_gz
- canu
- consensus
- digital_normalisation
- dsrc_to_gz
- falco
- fastp
- fastq_stats
- fastqc
- feature_counts
- freebayes
- freebayes_vcf_filter
- gz_to_bz2
- hmmbuild
- hmmscan
- index
- longorfs
- macs3
- makeblastdb
- mark_duplicates
- minimap2
- multiqc
- polypolish
- predict
- prokka
- quast
- rulegraph
- sambamba_filter
- sambamba_markdup
- samtools_depth
- sequana_coverage
- sequana_taxonomy
- snpeff
- snpeff_add_locus_in_fasta
- sort
- spades
- trinity
- trinity_quantify
- unicycler
- API reference
- Glossary